




Quickly compare and visualize distances between genomes, from assemblies (fasta) or reads (fastq), using Mash. Produces a distance matrix, trees (UPGMA, neighbour joining, minimum evolution) with optional bootstrap support, and an interactive PCoA plot.
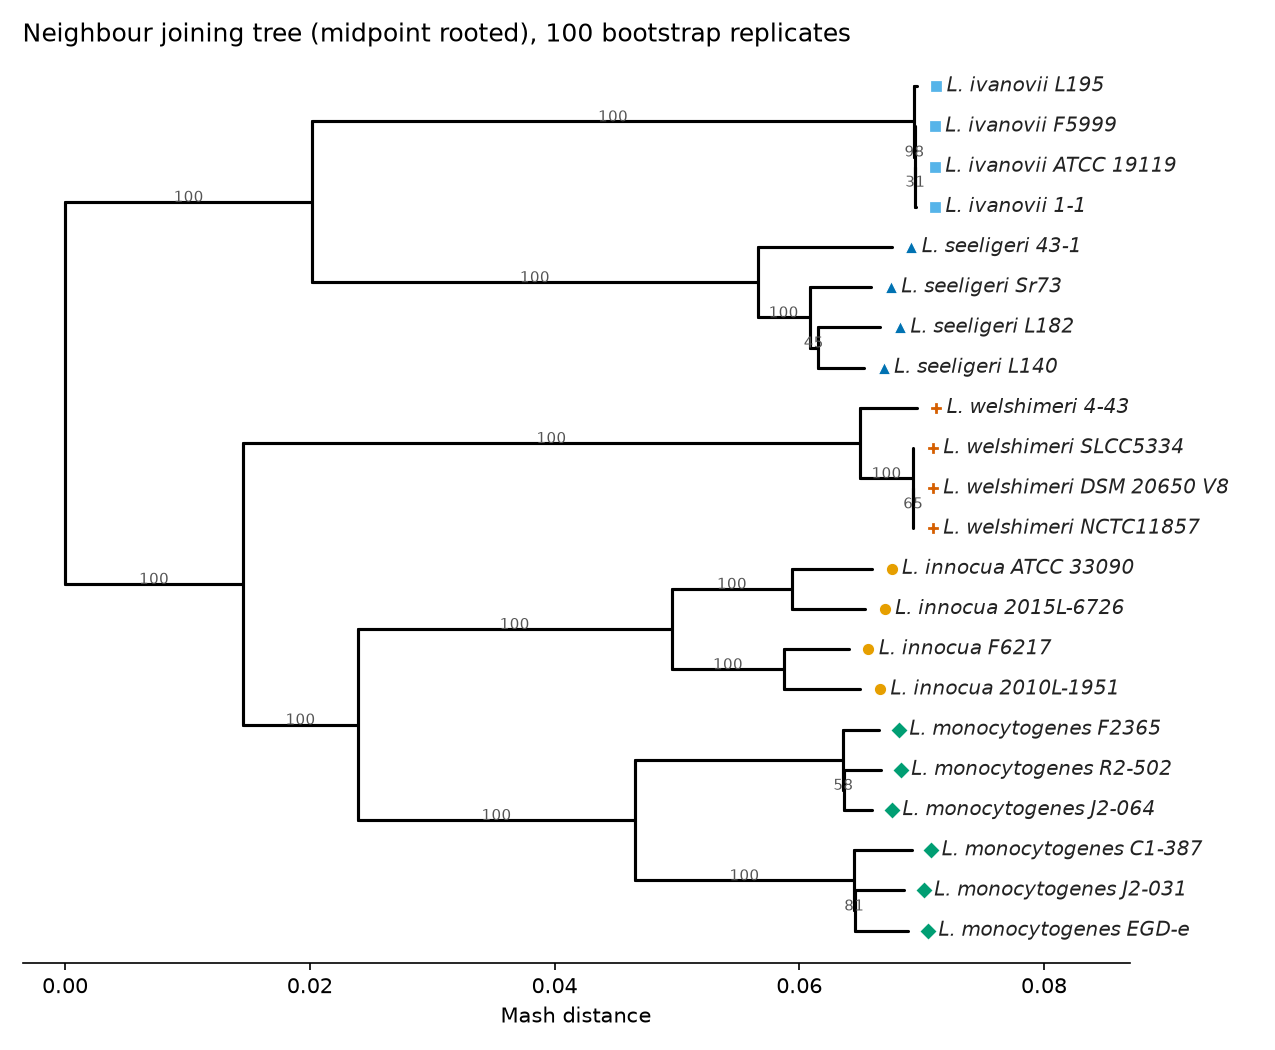
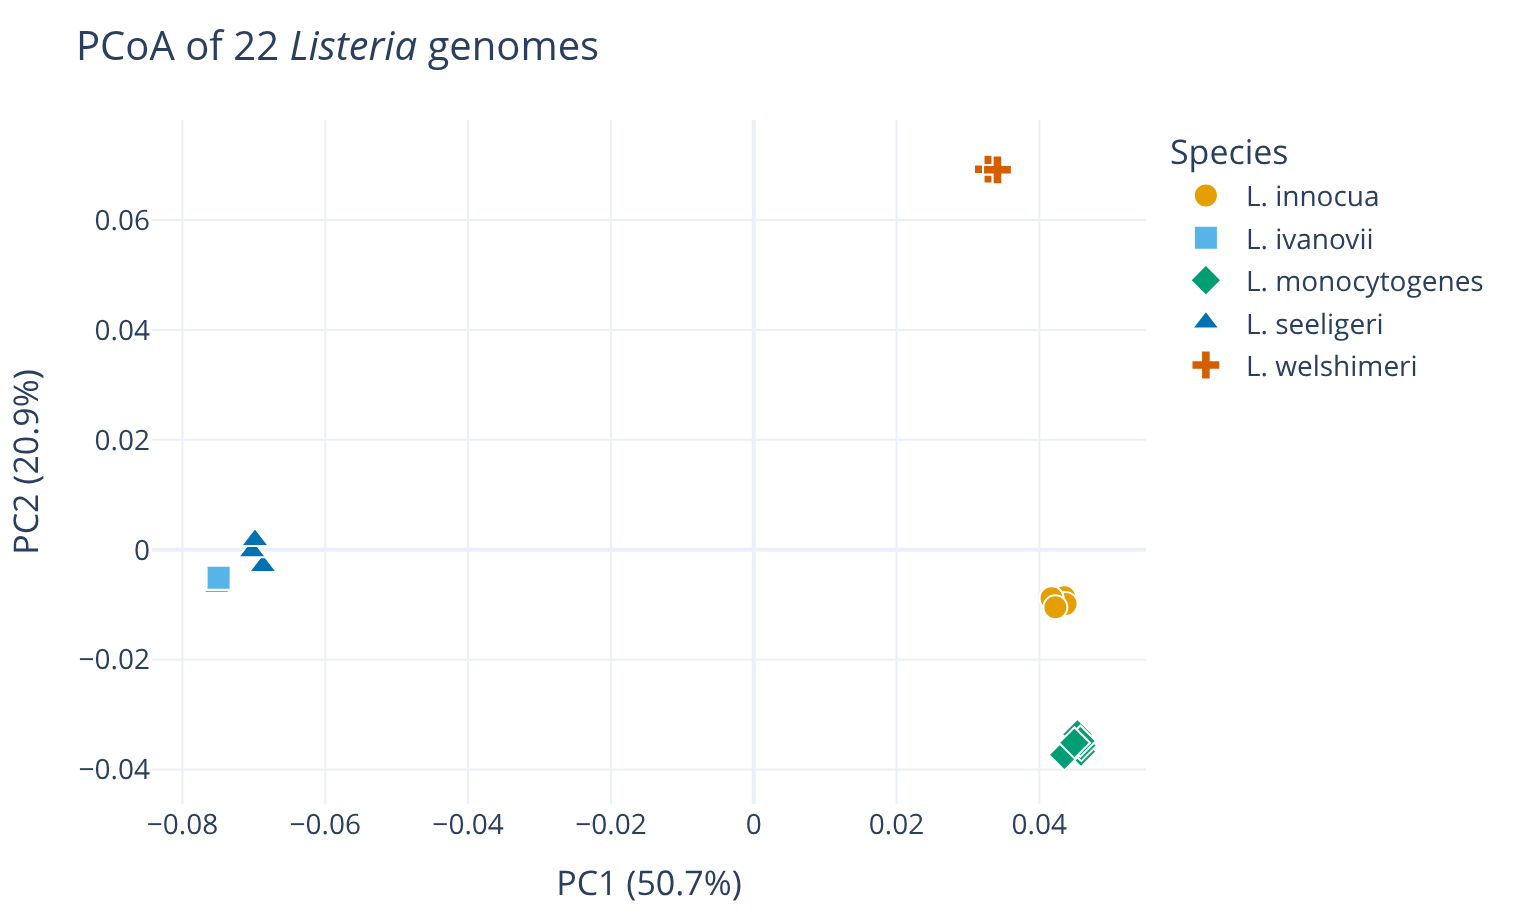
22 public Listeria genomes from the tutorial: 3 seconds, or under 2 minutes with 100 bootstrap replicates.
- Assemblies or reads: fasta or fastq (gzipped or not), paired-end reads combined per sample.
- Fast and scalable: thousands of genomes; sketches are computed in parallel, and adding genomes to an existing run only sketches them and measures their distances.
- Trees: UPGMA, neighbour joining and minimum evolution, with optional bootstrap support.
- Clusters at any distance thresholds, e.g. species (~95% ANI) or near-identical strains.
- iTOL annotations: colour strips for the metadata and clusters, to drag and drop on the trees.
- Interactive PCoA with metadata on hover and colourblind-friendly colouring by any metadata column.
- Safe by default: ambiguous sample names and unreadable files are reported, never silently merged or ignored.
flowchart LR
    A[fasta / fastq files] -->|mash sketch| B[one sketch per sample]
    B -->|mash triangle| C[pairwise distance matrix]
    C --> D[UPGMA / NJ / ME trees]
    C --> E[PCoA plot]
    B -. re-sketch with other hash seeds .-> F[bootstrap replicates]
    F -. support values .-> D
With pip. Mash is not available from PyPI: install it separately (e.g. conda install -c bioconda mash).
pip install genome-comparator
Or from source, in a conda environment that includes Mash:
git clone https://github.com/duceppemo/genome_comparator
cd genome_comparator
conda env create -f environment.yml
conda activate genome_comparator
pip install --no-deps .
genome-comparator -i /input/folder/ -o /output/folder/ --nj --pcoa
New to the tool? Follow the tutorial: it downloads 22 public genomes and walks through every output in a few minutes.
The wiki covers usage and options, output files, interpreting the results, bootstrap support, performance, related tools, troubleshooting and the FAQ.
If you use genome_comparator, please cite it and Mash, which computes the distances:
Duceppe M-O. genome_comparator: fast comparison and visualization of genome distances with Mash. Zenodo. https://doi.org/10.5281/zenodo.22920856
This DOI always points to the latest version; each release also has its own DOI, listed on Zenodo. GitHub's "Cite this repository" button gives the same citation in APA and BibTeX formats.
Ondov BD, Treangen TJ, Melsted P, Mallonee AB, Bergman NH, Koren S, Phillippy AM. Mash: fast genome and metagenome distance estimation using MinHash. Genome Biology 17, 132 (2016). https://doi.org/10.1186/s13059-016-0997-x
Bug reports, questions and pull requests are welcome: see CONTRIBUTING.md.
Marc-Olivier Duceppe, Canadian Food Inspection Agency (CFIA): marc-olivier.duceppe@inspection.gc.ca